Computes pairwise phi coefficients between binary categorical variables
with explicit 0/1 coding (1 == PositiveLevel from createBinaryMapping()),
then renders heatmap-style plots with raw and FDR-adjusted significance.
Usage
PlotPhiHeatmap(
  data,
  CatVars,
  Relabel = TRUE,
  binary_map = NULL,
  fdr_scope = c("matrix", "per_outcome", "per_predictor"),
  cluster_rows = FALSE,
  cluster_columns = FALSE,
  triangle = c("full", "upper", "lower"),
  IncludeTrendSymbol = FALSE,
  Data = lifecycle::deprecated()
)
Arguments
- data
A dataframe.
- CatVars
Character vector of binary categorical variable names.
- Relabel
Logical; if TRUE, uses sjlabelled variable labels for axes.
- binary_map
Optional mapping as returned by
createBinaryMapping(). If NULL, a mapping is created internally forCatVars.- fdr_scope
Either
"matrix"(default) or"per_outcome", passed toApplyFDRCorrection()."matrix"corrects across all p-values at once (historical behavior)."per_outcome"corrects separately within each y-axis variable (YVar); the Phi matrix is symmetric, so this treats each variable's row of tiles as one family.- cluster_rows, cluster_columns
Logical; cluster y-axis rows and/or x-axis columns using displayed Phi-coefficient profiles.
- triangle
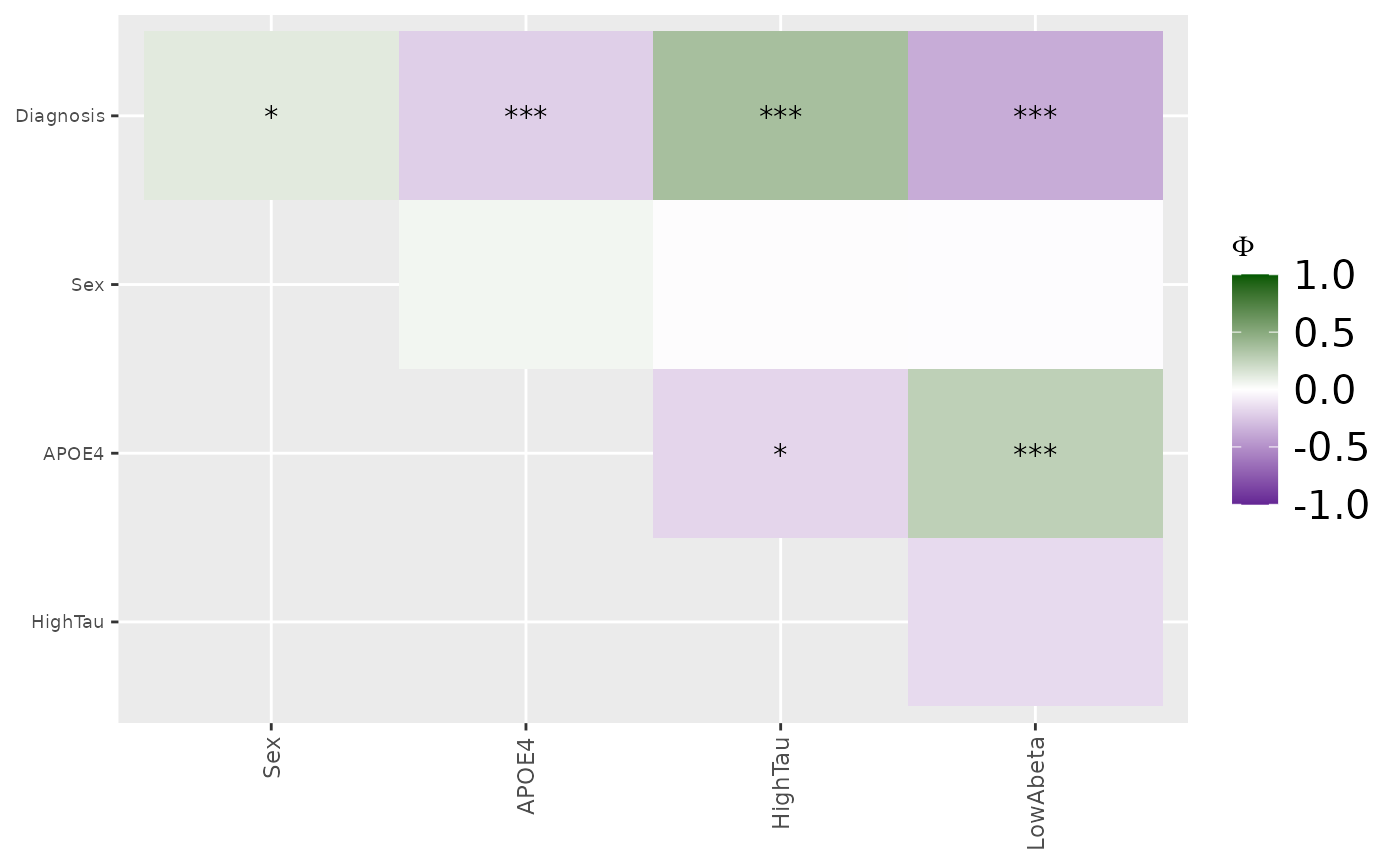
Display
"full"(default),"upper", or"lower"half of the symmetric Phi matrix. This affects the plot only; returned results remain complete.- IncludeTrendSymbol
Logical; add a plus sign for
0.05 < p <= 0.10.- Data
Deprecated (since 19.15.0). Use
datainstead.
Value
A list with:
Unadjusted: list(PvalTable, plot)FDRCorrected: list(PvalTable, plot)method= "Phi"RelabelBinaryMapping(used)
Examples
data(SampleData)
data(SampleVariableTypes)
Labelled <- RevalueData(SampleData, SampleVariableTypes)$RevaluedData
# CatVars must be binary (exactly two unique non-NA values). Derive a few
# more binary indicators so the matrix has off-diagonal structure to read;
# self-associations on the diagonal are masked out.
Labelled$APOE4 <- ifelse(
grepl("E4", as.character(Labelled$Genotype)), "Carrier", "Non-carrier")
Labelled$HighTau <- ifelse(
Labelled$tau > stats::median(Labelled$tau, na.rm = TRUE), "High", "Low")
Labelled$LowAbeta <- ifelse(
Labelled$Ab_42 < stats::median(Labelled$Ab_42, na.rm = TRUE), "Low", "High")
result <- PlotPhiHeatmap(
Labelled,
CatVars = c("Diagnosis", "sex", "APOE4", "HighTau", "LowAbeta")
)
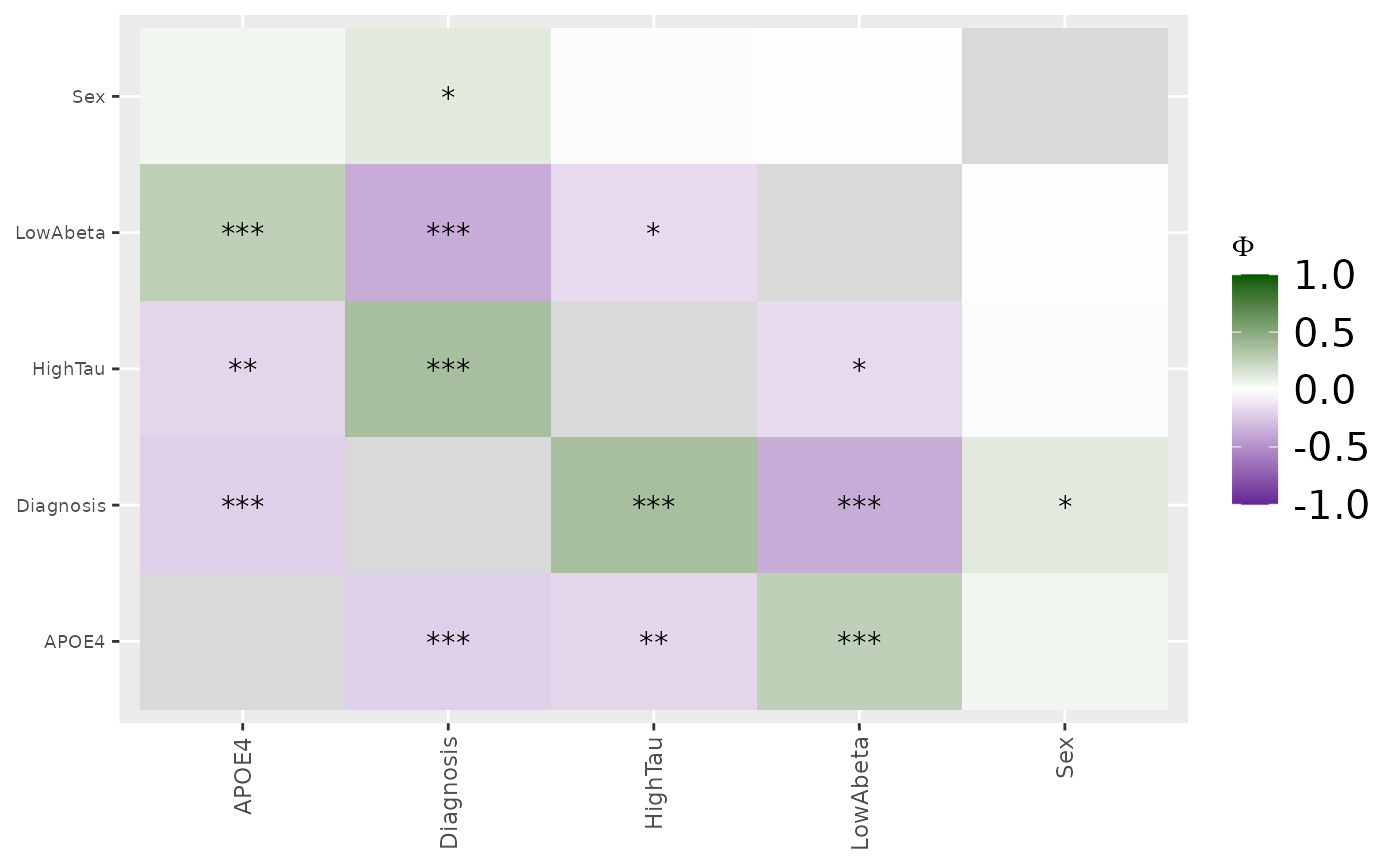
# Raw p-value phi heatmap
result$Unadjusted$plot
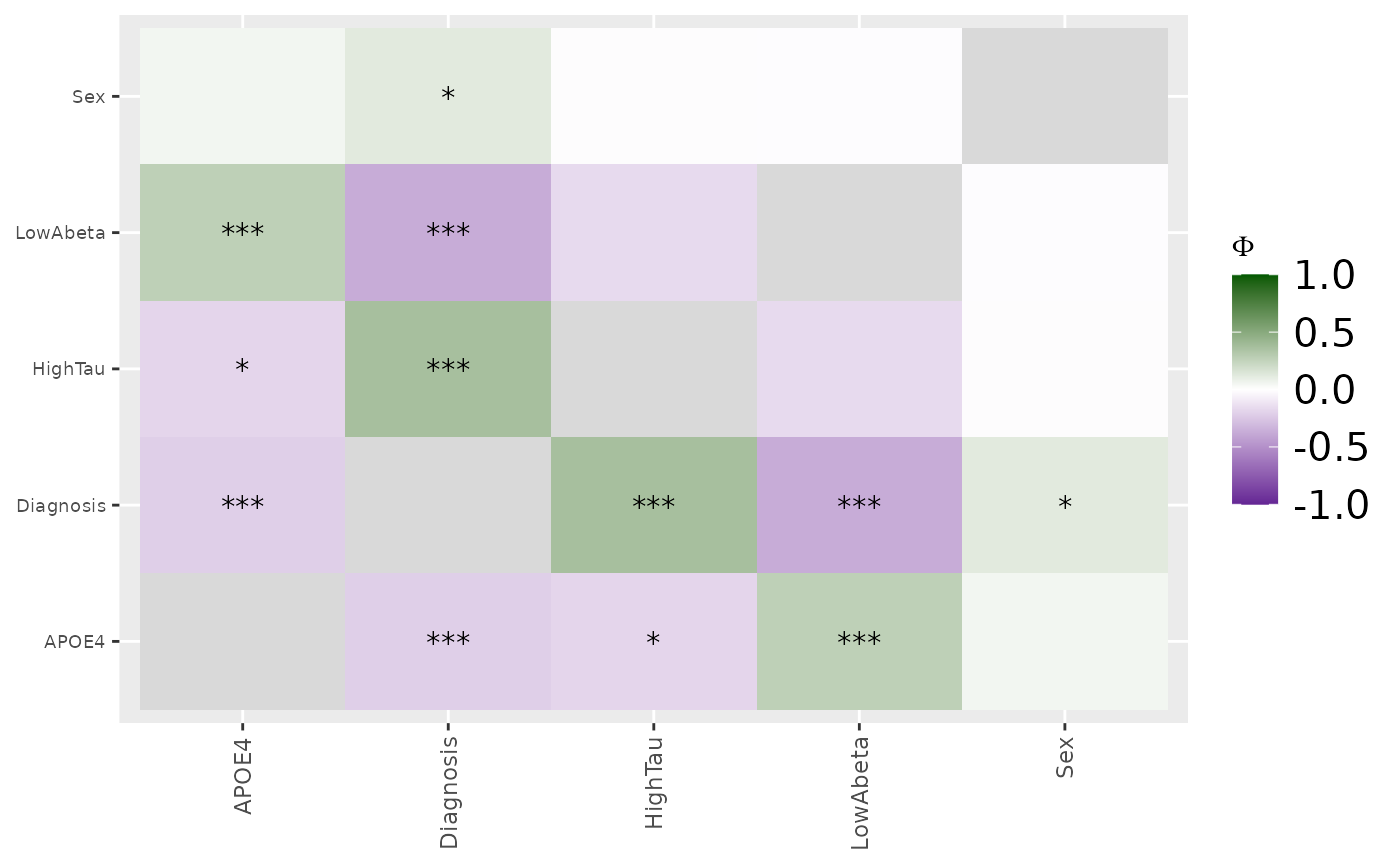
 # FDR-adjusted phi heatmap
result$FDRCorrected$plot
 # Show each binary-variable pair once.
upper <- PlotPhiHeatmap(
Labelled,
CatVars = c("Diagnosis", "sex", "APOE4", "HighTau", "LowAbeta"),
triangle = "upper"
)
upper$FDRCorrected$plot